How transcription factors (TFs) selectively recognize their genomic targets among numerous sites containing similar DNA motifs remains an open question. (1/n)
We evaluate the in silico mutations by testing their relative contribution to this count score. This is our ‘scale’ also in the context of cooperativity: the extent to which cooperative interaction terms contribute to or explain the variation in peak strengths.
Thanks, Anshul. Thank you for discussing this with us and for making the models and the data publicly available. It is really appreciated. We are looking forward to hearing more and comparing our approaches.
Great to see instant use of our resource. This is the advantage of fast open releases of data and model resources. But I disagree with the conclusions from this analysis. We have been working on a detailed analysis of these data & come to a very different conclusion. 1/
Our main interest is the differential binding strengths observed across peaks/ non-peaks, many of which do have the canonical motifs. So, in this analysis, we are only considering the variation in peak strength, as captured by the count score.
We believe that these findings highlight the need to critically assess prevailing assumptions and explore alternative mechanisms underlying selective TF binding.
Although detected in some cases, cooperativity had a negligible contribution to genome-wide binding predictions. Nevertheless, the models captured differences in binding preferences between TFs with similar motif preferences, even within the same cellular environment.
We believe that these findings highlight the need to critically assess prevailing assumptions and explore alternative mechanisms underlying selective TF binding.
Although detected in some cases, cooperativity had a negligible contribution to genome-wide binding predictions. Nevertheless, the models captured differences in binding preferences between TFs with similar motif preferences, even within the same cellular environment.
(7/7) This is the lab’s first mammalian study, extending principles of IDR-directed genome targeting from yeast to mammalian transcription factors while introducing a mammalian ChEC-seq platform for high-resolution mapping of protein-DNA interactions. https://t.co/IOTVuioVjY
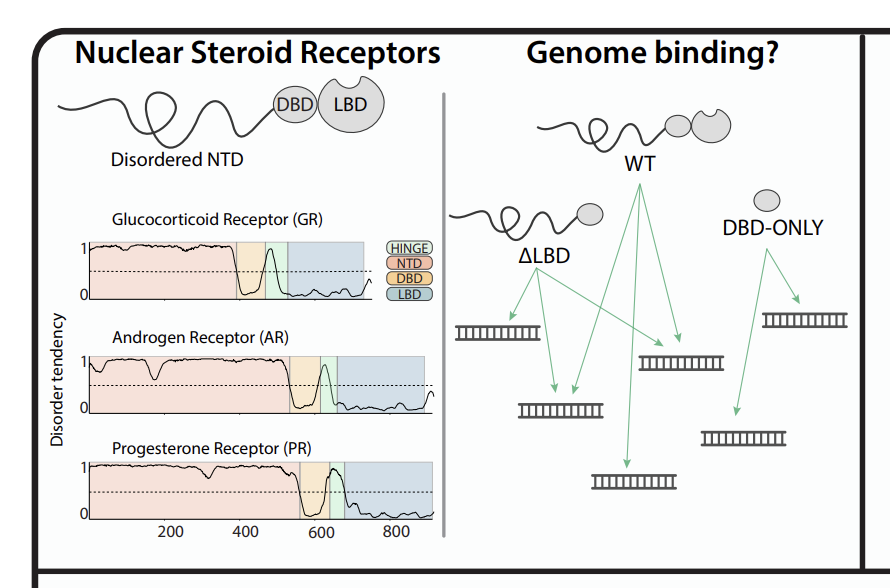
(1/7) How do steroid receptors identify their target sites within the mammalian genome, despite the abundance of similar DNA motifs? We tackled this question by adapting ChEC-seq for mammalian tissue culture, enabling footprint-resolution mapping of transcription factor binding.
(6/7) The NTDs also direct genome targeting to distinct SR motifs, but not to other Transcription factor’s motifs in budding yeast, where mammalian cofactors are absent.
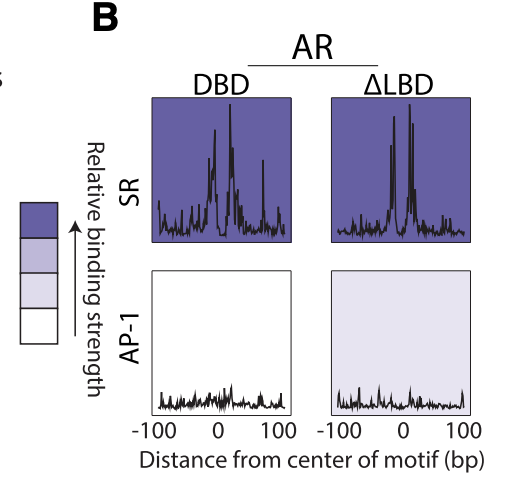
(5/7) Footprint analysis uncovered an unexpected role for AP-1. Rather than simply recruiting steroid receptors, AP-1 strongly shapes binding when the NTD is absent. The NTD relieves this AP-1 bias, enabling receptors to efficiently bind canonical steroid response elements.