Single-base methylation across the whole genome: WGBS with >99% conversion efficiency, ~30X coverage, and full DMR analysis. For cfDNA/FFPE, we'll steer you to EM-seq instead.
https://t.co/25hwfjXeHu
#Epigenetics#WGBS
Maize's selfish chromosomes cheat Mendel. WGBS by CD Genomics helped Brady & Dawe (PLOS Genetics 2025) show each drive haplotype carries a distinct methylation signature.
https://t.co/ihB0Q6EIgW
#Epigenetics#Genetics
Methylation, RNA modifications, chromatin — one platform instead of stitched-together vendors. CD Genomics epigenomics sequencing covers WGBS to CUT&Tag under one QC framework.
https://t.co/pNqXN7JlaI
#Epigenomics#Genomics
7,284 nanopore methylomes: an aging clock at ~2.4-yr error — and parent-of-origin-specific methylation drift at imprinted loci only long reads can phase (Sigurpalsdottir et al., Nat Commun 2026).
https://t.co/7SgWnuCsKf
#Epigenetics#Nanopore
EBV-positive DLBCL tumors carry a measurably distinct methylome — mapped by DNA methylation array services from CD Genomics (Volaric et al., Cancers 2025, CC BY).
https://t.co/m6YXsY97g9
#Epigenomics#CaseStudy
No bisulfite. Single molecules. Phased methylation.
Long-read DNA/RNA methylation, Fiber-seq & custom workflows — direct 5mC/5hmC calls with haplotype resolution.
https://t.co/7gdAuUTcFj
#Epigenetics#LongReadSequencing
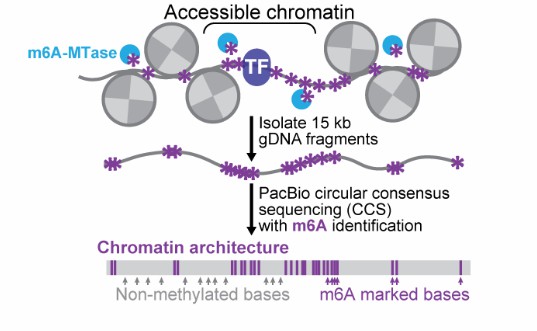
Accessibility + haplotype + nucleosomes, from one library. Fiber-seq: m6A-labeled open chromatin read by PacBio HiFi, single-molecule resolution.
https://t.co/eCySgaPF1c
#FiberSeq#LongReadSequencing#Epigenomics
Bulk m6A is an average over millions of cells. m6A-ARTR-DBiT reads the mark in situ on tissue sections and barcodes cDNA spatially — ~20 µm pixels, regional m6A maps in mouse brain.
https://t.co/BKsXnu2ojY #m6A#SpatialOmics
FIONA1 deposits m6A at the 3' end of FLC. That mark stabilises the mRNA; unmethylated FLC is degraded fast, and flowering time shifts.
mRNA-seq + Nanopore direct RNA-seq (CD Genomics) + MeRIP-seq. https://t.co/J6ldITmxMr #m6A
Most m5C in a total RNA prep sits on tRNA and rRNA. If your claim is about mRNA, depletion has to show in the mapping stats.
Bisulfite, oxBS or RIP — the assay decides what you can see. https://t.co/gTp6Ie0qAW #m5C#Epitranscriptomics
m6A-seq locates m6A within ~100 nt, not at a base. Fragment size sets the resolution ceiling — a peak is a window, not a coordinate.
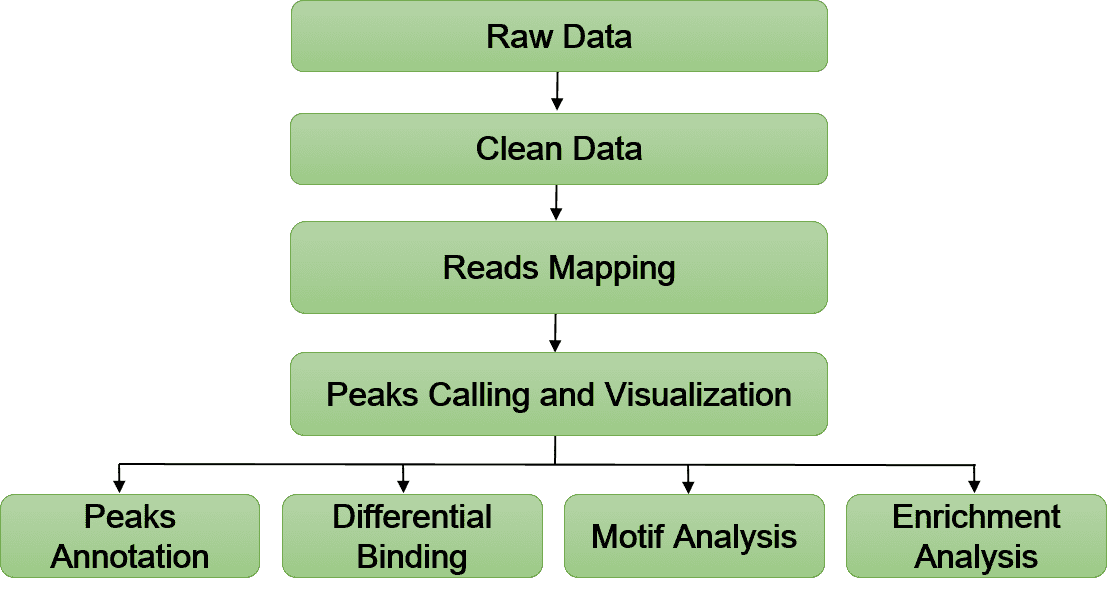
Peak calling, DRACH QC, differential methylation. https://t.co/0SuyVa3J2N #m6A
Customer case: the CLK2/SRSF9 splicing axis drives AR-V7 in prostate cancer. Van Goubergen et al. (Mol Oncol 2025) used nanopore sequencing provided by CD Genomics for isoform-resolved transcripts. https://t.co/cqUTc8sAct #LongReadSequencing#CancerGenomics
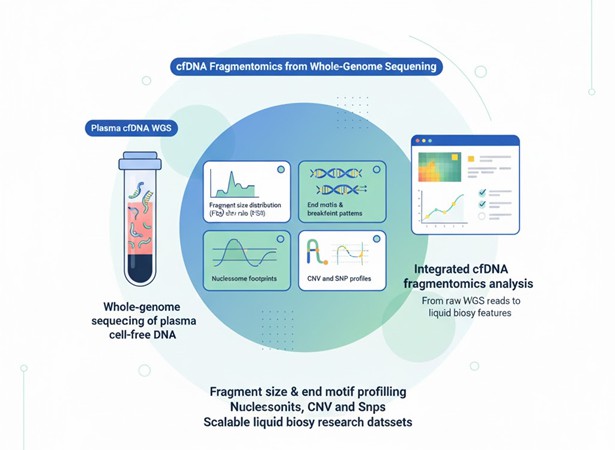
2 mL plasma, 1–5x WGS. cfDNA fragmentomics reads fragment sizes, end motifs, nucleosome footprints, CNV — a tissue-origin signal beyond mutations.
Full service + pipeline: https://t.co/0CBz2obaI5
#LiquidBiopsy#cfDNA#NGS
11,692 genes silenced. 2.5M single cells profiled.
A genome-scale CRISPRi atlas of human iPSCs (Nat Biotechnol, 2026) maps how genotype shapes cell state — data and pipelines fully open.
https://t.co/79RxRyyMyG
#CRISPRi#iPSC
A genome-wide CRISPR screen in haploid hESCs revealed ATF7IP as a suppressor of paternal imprinting (Nat Commun).
RRBS library prep & sequencing for the methylation maps: provided by CD Genomics.
https://t.co/UiBL5DMOqP
#CRISPR#Epigenetics
The RNA you want may not be in cells — it's in transit.
Exosomal RNA sequencing: miRNA, mRNA & lncRNA from serum, plasma, urine, or culture supernatant.
https://t.co/SoLntKUs4C
#Exosomes#LiquidBiopsy
Pooled CRISPR screens live or die on the readout.
CRISPR screen sequencing + MAGeCK analysis: sgRNA abundance, fold changes, ranked gene hits from 5×10⁶ cells or ≥1.5 μg gDNA.
https://t.co/X38qvBlH92
#CRISPR