@picoplankton I think the reverse primers in two different papers (Paul-2000-Marine ecology progress series & Wawrik-2004-QUATIC MICROBIAL ECOLOGY) in your lab were different. Could u give me some suggests about how to use them, thanks so much.
Dear Dr. Malik @_biogeek I am reading your paper "Land use Driven change in soil pH affects microbial carbon cycling processes", in which there was a method called "slope failure test", and could you show that how you decide the intervals and the units of pH, please?
Dear @hmm3r I used the command hmmbulid (Hmmer3) build a hmm model from a seed txt that download from Pfam. And then i copied it under genewise to compared a DNA with it, tbut after run that, it said "Can not open hmm file". Could u help me to solve this problems?
@ewanbirney Dear professor @ewanbirney, i just want to thank you for your help. I have installed wise2.2.0 successfully. Because I have few knowledge about linux, and I dropped wise2.4.1 and tried wise2.2.0. Then, it succeed. Thanks so much for your kindly help.
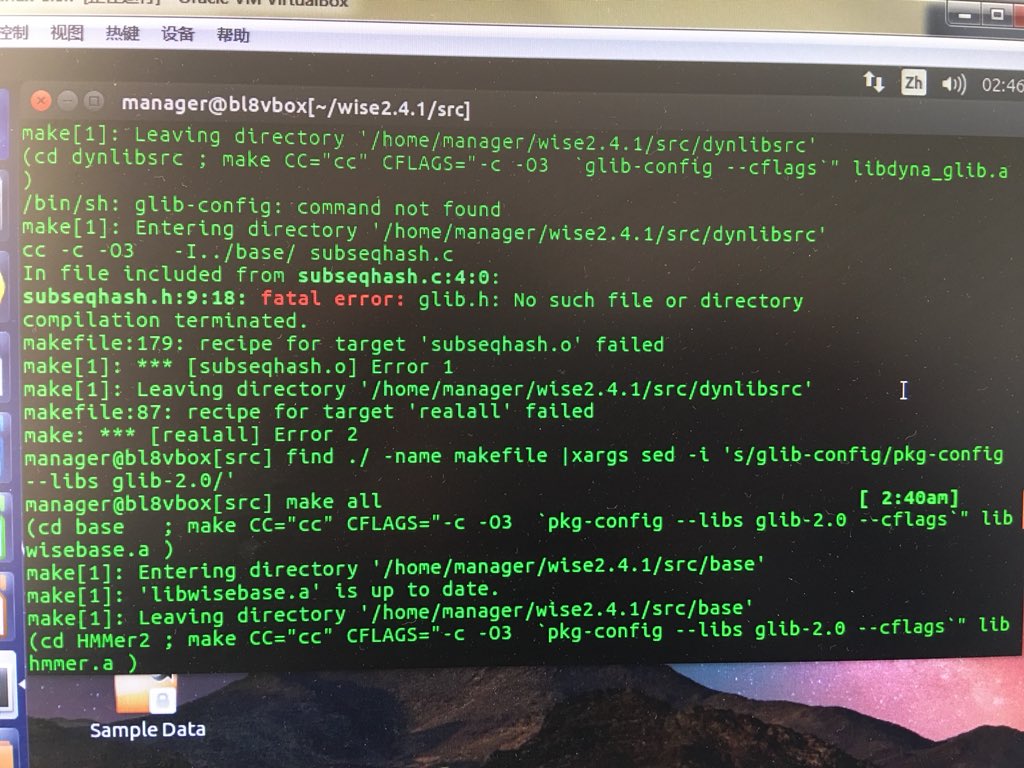
@ewanbirney Dear profession, I changed all the getline into wise_getline in "src", and then it showed that "subseqhash.h:9:18", and then i use "find ./ -name makefile |xargs sed-i 's/glib-config/pkg-config--libs glib-2.0/'" ,which from web. Then, it another wrong accured. Could u help me
@ewanbirney Thanks for your suggestion, in fact, i need use the HMMs to stduy the Family: Glyco_hydro_7(PF00840). I will try again according to your suggestion, and thanks so much for your reply.
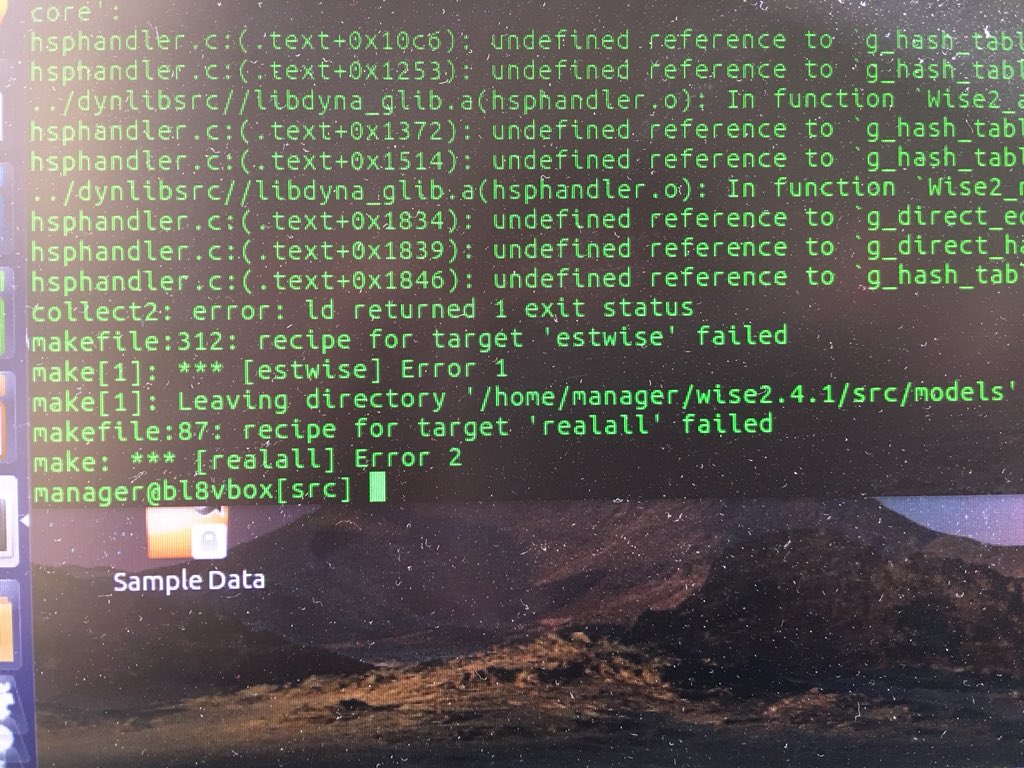
Dear professor @ewanbirney , i had a problem during the install of genewise2.4.1. The picture is after " make all"in "src". Could you give me some suggestion? I'm a student from CAS, China. I'm a new one in bioinformation, and really hope for your guide.